1. Why HMO production is a hard glycobiology problem
2. Precision fermentation: The engineering framework for scalable HMO production
3. Scale-up: From shake flask to industrial fermentor
4. Emerging frontiers: Engineering beyond the approved five HMOs
5. Analytical quality control: Verifying what you have made
6. Frequently asked questions
HMO manufacturing is difficult because the target molecules are not linear commodity sugars. They are structurally precise glycans with activity tied to linkage, branching, terminal decoration, and stereochemistry. A small change in fucosylation or sialylation can alter receptor binding, microbiome selectivity, and immune interaction. That structural sensitivity makes HMO production a well-known glycobiology bottleneck.
Precision fermentation offers a scalable route. Microbial hosts can be engineered to generate nucleotide-sugar donors, express pathway-specific glycosyltransferases, and assemble defined structures in a controlled fermentation process. In practice, producing HMOs through microbial synthesis shifts the core challenge from step by step chemical synthesis to metabolic engineering.
For a full overview of HMO structures, types, and commercial applications, see our complete guide to human milk oligosaccharides.
More than 200 HMO structures arise from combinations of five main monosaccharides: glucose, galactose, N-acetylglucosamine, fucose, and sialic acid. The difficulty sits in how these units are connected. HMOs contain α and β glycosidic linkages across 1→2, 1→3, 1→4, and 1→6 positions, often within branched architectures.
Each linkage type requires tight control over enzyme specificity, donor availability, acceptor recognition, and reaction order. Fucosyltransferases, sialyltransferases, galactosyltransferases, and N-acetylglucosaminyltransferases do not act as interchangeable tools. Their selectivity defines the final glycan.
Chirality adds another constraint to chemical synthesis routes. Stereocontrol is not optional in HMO synthesis. Chemical routes must manage anomeric configuration and regioselectivity through protecting group strategies, leaving groups, catalysts, and purification at each stage. With every additional monosaccharide, the synthetic route expands sharply. Step count rises. Yield compresses. Analytical burden increases. Enzymatic and microbial routes largely sidestep these challenges, since glycosyltransferase enzymes are inherently stereospecific, installing each sugar residue with the correct anomeric configuration and regiochemistry. This is one of the fundamental practical advantages of biological production over chemical synthesis of HMOs.
Human milk extraction was the original source of HMOs, but it cannot support industrial supply. Ethical constraints, limited donor material, batch variability, and regulatory complexity make it unsuitable for broad commercial production.
Chemical synthesis can produce defined HMOs, especially for standards and research materials. Scale is the challenge. Protecting group chemistry reduces step economy, solvent burden is high, and stereochemical control becomes harder as structures become larger and more branched.
Enzymatic synthesis improves selectivity. Even so, throughput can be limited by enzyme cost, nucleotide-sugar substrate supply, cofactor recycling, and downstream purification. Precision fermentation addresses these constraints by internalising parts of donor generation and glycan assembly inside microbial production systems.
Precision fermentation reframes HMO manufacturing as a chassis, pathway, and process optimisation challenge. The objective is controlled assembly of a defined glycan from lactose and activated nucleotide-sugar donors, with the correct linkage pattern and impurity profile.
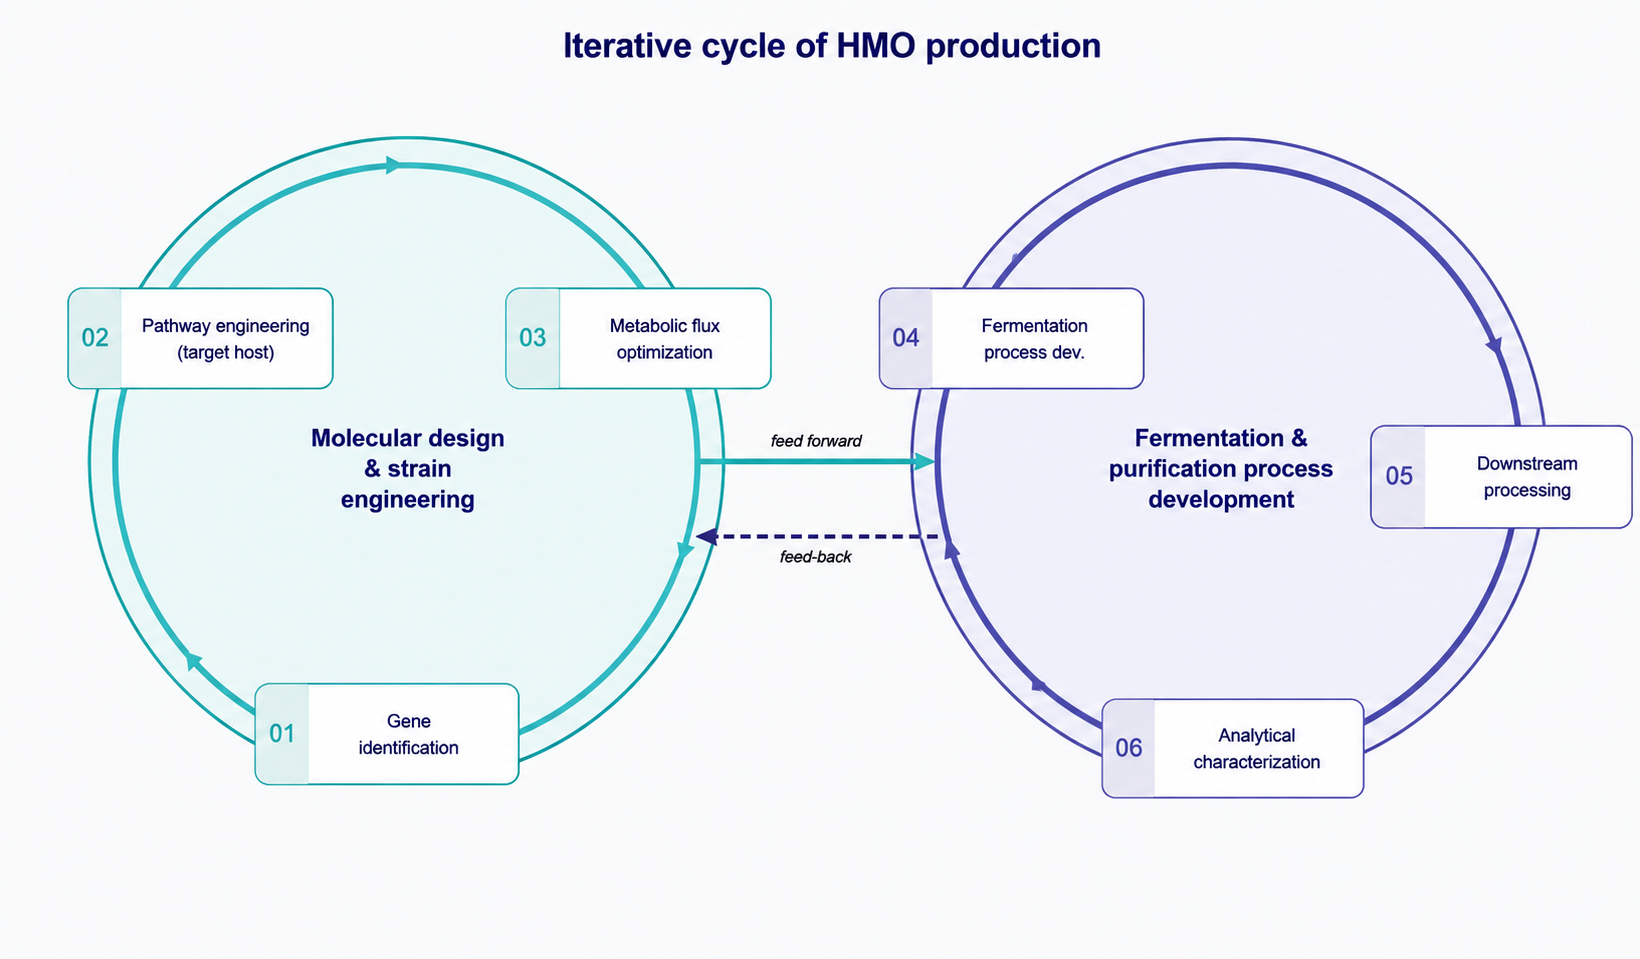
Development of a generic precision fermentation workflow for HMOs usually follows this logic:
1. Gene identification
Start with the target HMO. Look for glycosyltransferase genes that fit. Donor preference, acceptor specificity, linkage type, and host compatibility all matter.
2. Pathway engineering
Insert the selected glycosyltransferase genes into the target host (typically E. coli).
Remove or block competing pathways. For lactose‑based routes, disabling lactose
catabolism genes like lacZ or lacY may help prevent acceptor loss or unwanted transport effects.
3. Metabolic flux optimisation
Boost the supply of precursor building blocks. Key donors include UDP‑galactose,
GDP‑fucose, and CMP‑Neu5Ac. Which ones matter depends on whether the target is neutral, fucosylated, or sialylated.
4. Fermentation process development
Fine‑tune fed‑batch conditions. Set the feeding strategy, balance carbon sources, control dissolved oxygen and pH, time any induction steps carefully, and manage by‑products.
5. Downstream processing
Get the HMO out and clean it up. Steps may include harvesting, filtration,
chromatography, activated carbon, crystallisation, or spray drying. The optimal sequence depends on product localisation, impurity profile, and final purity requirements.
6. Analytical characterisation
Check identity, purity, and safety. Common methods include HPLC, LC‑MS, NMR for structure, assays for residual host‑cell impurities, and microbiology and endotoxin testing when needed.
Feature | E. coli | Yeast |
Genetic toolkit | Highly mature. Efficient knock-ins, knock-outs, plasmid systems, genomic integration, and pathway refactoring | Mature toolkit, strong integration systems, stable expression options, and broad industrial fermentation experience |
Growth rate | Fast growth and short iteration cycles | Slower than E. coli, but robust at industrial fermentation scale |
Endotoxin | Lipopolysaccharide endotoxin requires specific downstream control and testing | No LPS endotoxin, which can simplify some impurity considerations |
Regulatory status | Strong precedent for microbial fermentation products, including food and ingredient applications | Strong industrial precedent across food, enzymes, proteins, and speciality ingredients |
Glycosylation | Well‑established production host for all types of HMOs | Native glycosylation machinery available |
Commercial examples | Generic commercial precedent exists for bacterial fermentation of selected HMOs | Generic commercial precedent exists for yeast-based production of food and speciality biochemicals |
The practical question is usually not whether E. coli or yeast is “better.” It is whether the host can maintain flux through the desired HMO biosynthesis pathway while tolerating precursor drain, membrane transport stress, osmotic pressure, and product-associated burden.
The glycosyltransferase is the central specificity element in precision fermentation HMO production. GTs catalyse transfer of sugar moieties from activated nucleotide-sugar donors to acceptor substrates. In HMO pathways, lactose often acts as the starting acceptor. Donors may include UDP-Gal, GDP-Fuc, or CMP-Neu5Ac, depending on the target structure.
A glycosyltransferase HMO strategy depends on linkage control. A fucosyltransferase forming an α1→2 linkage does not substitute for one forming α1→3 or α1→4. The same principle applies to sialyltransferases and galactosyltransferases. Donor preference, acceptor tolerance, catalytic efficiency, regioselectivity, and expression behaviour all matter.
Many GTs contain conserved catalytic features, including the DxD motif in metal-dependent enzymes. Structural families such as GT-A and GT-B folds provide useful engineering context, although fold class alone does not predict industrial performance. Local active-site geometry, acceptor binding, donor positioning, and protein stability influence final pathway output.
With a specific target carbohydrate in mind, the process typically starts by exploring a wealth of sequence information in public and proprietary databases. Using specialised bioinformatic algorithms, a few dozen to hundreds of candidate enzyme sequences are selected for synthesis and screened for the desired glycosyltransferase activity using high-throughput screening assays in the lab. The best-performing hits are subsequently characterised and verified in depth.
To further optimise the selected glycosyltransferase enzymes, two broad engineering strategies are common. Rational design uses sequence, structural, and mechanistic information to modify residues linked to substrate recognition, catalytic efficiency, or stability. Directed evolution screens mutant libraries for improved activity, altered specificity, or better expression in the selected chassis. In practice, both approaches can be combined. Rational design narrows the search space. Directed evolution captures functional improvements that are not obvious from structure alone.
GT engineering also has a systems dimension. A highly active enzyme can create imbalance if donor supply is limiting. A slow enzyme can cause intermediate accumulation. A broad-specificity enzyme can generate side products. For HMO strain engineering, enzyme selection is therefore coupled to flux, transport, product export, and purification burden.
Precursor supply often limits HMO pathway performance before the terminal GT step does. For fucosylated HMOs, GDP-fucose availability is central. The de novo route converts central carbon intermediates into GDP-fucose through enzymes such as Gmd and WcaG in E. coli contexts. Alternative designs may use salvage logic, depending on substrate availability and host capability. Upstream mannose-related steps involving ManB and ManC can also influence donor formation.
For sialylated HMOs, CMP-Neu5Ac supply becomes the key engineering target. Overexpression of genes such as neuB and neuA can support Neu5Ac synthesis and activation, provided the pathway has sufficient precursor input and does not impose excessive metabolic burden. Sialylated products often add further analytical and purification complexity because charge state, isomer profile, and related acidic impurities require tight control.
Lactose handling is another design constraint. The cell must maintain intracellular lactose as an acceptor while avoiding catabolism. In bacterial systems, this can involve control of lactose uptake and hydrolysis through genes such as lacZ and lacY. The aim is not simply high lactose concentration. The aim is acceptor availability at the right point in the pathway, without growth inhibition, excessive overflow metabolism, or unwanted by-product formation.
Effective HMO strain engineering therefore balances three layers at once: donor biosynthesis, acceptor management, and GT-catalysed assembly. Fermentation development then tests whether that engineered architecture remains stable under process conditions. Fed-batch feeding, oxygen transfer, pH control, and harvest timing all influence the final product profile.
HMO scale-up fermentation is not a linear enlargement of a shake-flask protocol. The biology changes with oxygen transfer, mixing time, feed gradients, heat removal, foam behaviour, and cell-state heterogeneity. A strain that performs cleanly at millilitre scale can show different flux distribution in a production fermentor, especially when precursor demand, lactose availability, and by-product formation start to compete.
For HMOs, scale-up is usually about maintaining the same biosynthetic logic under industrial constraints. The target remains the assembly of defined glycans. The harder task is keeping productivity, impurity profile, and process robustness aligned as volume increases.
Batch fermentation is useful for early screening and baseline physiology. In batch mode, all nutrients, including the carbon source and lactose acceptor, are loaded at the start of the run, with no further additions until the process ends. It gives a controlled view of growth, substrate consumption, and product formation without complex feeding logic. For commercial HMO production, however, simple batch mode usually provides limited control over lactose availability, carbon flux, and metabolic burden.
Fed-batch is the industry standard for many HMO processes. Unlike batch mode, fed-batch adds nutrients incrementally over the course of the run, allowing e.g. the carbon source to be adjusted as the culture grows. The carbon feeding strategy is especially important. A sufficient amount of carbon source is essential, but excess can affect osmotic pressure, transport behaviour, residual sugar load, and downstream burden.
Nitrogen balance and pH control also shape the final product profile. At high-cell-density scale, two physical constraints dominate: oxygen transfer and CO2 removal. Oxygen transfer rates become more difficult at scale. High-cell-density cultures increase oxygen demand, and insufficient oxygen transfer could shift metabolism toward unwanted by-products. CO2 removal also becomes critical. Accumulated CO2 can affect pH, growth physiology, and enzyme performance.
Continuous fermentation is technically attractive for productivity and asset utilisation, with cells and nutrients continuously fed in and product-containing broth continuously withdrawn to maintain a steady state, but it adds its own control problems. Genetic stability, contamination risk, steady-state impurity drift, and long-duration process validation become major considerations. For HMOs, fed-batch often remains the practical middle ground between control, robustness, and regulatory familiarity.
HMO purification during downstream processing depends on host organism, product localisation, broth composition, product charge, residual lactose, salts, host-cell impurities, and final purity requirements. There is no single universal purification solution.
Early recovery may include cell removal by centrifugation or microfiltration. If the product is intracellular or partly cell-associated, harvest logic can differ from a fully extracellular process. Clarified streams then require impurity reduction. Options include activated carbon treatment for decolourisation, ion exchange chromatography for charged impurities, desalting, and removal of host-derived contaminants.
Concentration steps may use nanofiltration or ultrafiltration. The choice depends on molecular size, salt profile, and what the process needs to achieve. Nanofiltration can help concentrate target HMOs and cut down low‑molecular‑weight impurities. Ultrafiltration may help remove larger impurities or condition the process stream.
Final polishing depends on the molecule and the target specification. Neutral HMOs and acidic HMOs behave differently during purification. Charge, solubility, and co‑purifying species all play a role. Possible finishing routes include crystallisation or spray drying to produce a powder. Which route to pick depends on product stability, purity targets, moisture limits, particle properties, formulation needs, and regulatory expectations.
Analytical support is not an afterthought in downstream development. HPLC, LC-MS, NMR, residual protein assays, residual DNA assays, bioburden and endotoxin testing where applicable help define whether purification is removing the right impurities, not merely concentrating the target glycan.
The obvious cost inputs are feedstock, utilities, fermentation time, and downstream consumables. The hidden costs sit in productivity, impurity control, and yield recovery across unit operations.
Lactose and sucrose are key feedstocks, but feedstock cost is just one component of the broader process economics. Strain productivity has a larger effect because titre, rate, and yield determine fermentor occupancy and downstream load. A process with higher product concentration can reduce water handling, purification volume, and drying burden.
Scale also matters. Larger fermentors reduce certain fixed costs per kilogram, yet they amplify oxygen transfer limits, mixing gradients, foam control, and heat removal. Downstream processing can dominate the cost of goods if impurity profiles are complex or product recovery is low.
For HMO scale-up fermentation, the lowest-cost process is not simply the highest-titre process. It is the process that balances strain performance, feed strategy, purification efficiency, analytical control, and final product quality.
The first commercial HMO wave focused on structures with tractable pathways, clear demand, and stronger regulatory momentum. That made sense. Molecules such as 2′FL became useful case studies for 2’FL fermentation production because the pathway is comparatively compact and industrially learnable. The next frontier is different. More complex HMOs require longer biosynthetic logic, tighter enzyme ordering, and cleaner control of intermediate pools.
More complex structures such as LNFP and LNDFH push beyond single-decoration pathways. Multiple glycosyltransferases must act in sequence, often with narrow acceptor preferences and competing side-product risks. The pathway is no longer just donor supply plus one terminal transfer. It becomes an enzyme cascade challenge.
DSLNT is a relevant example because nobody produces it commercially yet. Its biological interest is clear due to its potential role in reducing NEC risk (2). However synthesis remains challenging and researchers are searching for efficient sialyltransferase enzymes to allow industrial manufacturing. In addition, the molecule requires coordinated assembly of a larger scaffold with specific sialylation and branching logic.
Note that for certain target structures, a plausible synthesis route may combine fermentation-derived core HMO scaffolds with enzymatic elaboration steps. Microbial production can deliver the backbone or intermediate. Enzymatic synthesis can then add difficult terminal decorations under more controlled conditions.
Engineered crops are also entering the HMO discussion. A 2024 Nature Food study reported plant-based production of diverse HMOs by using plant carbohydrate metabolism as a photosynthetic production platform (1). The same work included complex structures such as lacto-N-fucopentaose I, which makes it relevant to the next stage of HMO biomanufacturing.
The appeal is obvious. Plants offer low-cost biomass, solar-driven carbon capture, and agricultural scale. The open questions are equally large. Regulatory frameworks for food-grade transgenic production remain complex. Extraction must be efficient. Product consistency has to meet ingredient specifications. Field variability, tissue-specific accumulation, and co-extracted plant metabolites all become process variables.
Computational design is becoming more useful for HMO strain engineering. Constraint-based metabolic modelling can prioritise pathway designs before wet-lab screening. Such models help evaluate flux distributions, knock-out logic, and precursor drain, and can support pathway search and host-route comparison.
AI-guided directed evolution may also improve glycosyltransferase HMO performance. Models can rank mutations for activity, stability, donor preference, or altered acceptor scope. Digital twin fermentation adds another layer by linking feed profiles, oxygen transfer, pH, growth state, and product formation. That matters when small pathway changes create large process effects. Recent literature highlights the role of in silico metabolic modelling in pathway design.
HMO production does not end at titre or recovery. In HMO glycan manufacturing, the critical question is whether the isolated material is the intended structure, at the required purity, with process-related impurities under control. A supplier dossier should therefore include orthogonal identity data, not only a chromatographic purity value.
HPLC can establish purity and separate related carbohydrates, residual lactose, side products, and process impurities. LC-MS confirms molecular mass and helps detect isobaric or near-mass contaminants when method resolution is adequate. NMR remains the strongest structural confirmation tool because it verifies linkage position, anomeric configuration, and substitution pattern. For HMOs, that distinction matters. A correct molecular mass alone does not prove the correct glycosidic architecture.
Process safety data should sit beside structural data. Residual protein, host-cell DNA, endotoxin where applicable, heavy metals, and microbiological absence all affect suitability for regulated or formulation-sensitive applications. Powder specifications also need moisture or water activity, since hygroscopic behaviour can affect stability, flow, and storage performance.
Parameter | What it confirms | Method |
HMO identity and purity (% HMO) | Confirmation of HMO identity and free from impurities and by-products | HPLC, MS, NMR |
Residual protein | Removal of fermentation host proteins | BCA / Bradford assay |
Endotoxin | Safety-critical for E. coli-derived products | LAL assay |
Residual DNA | Process safety for regulated applications | qPCR |
Moisture / water activity | Stability of powder form | Karl Fischer / water activity meter |
Appearance | Visual and physical characterisation | Visual inspection / spectrophotometry |
Heavy metals | Safety — absence of toxic metal contaminants | ICP-MS |
Microbiology | Safety-critical; absence of microbial contamination | Plate count / compendial methods |
A robust QC package should also show method suitability, batch consistency, and impurity tracking across development lots. For novel or less common HMOs, reference standard quality becomes especially important. Without defensible structural assignment and impurity profiling, biological testing results can become difficult to interpret.
Interested in the health effects of HMOs rather than their production? Our guide to HMOs for gut health, immunity, and beyond covers the mechanisms, clinical evidence, and applications in detail.
Precision fermentation uses engineered microbial hosts as cell factories to produce specific, high-value compounds such as proteins, lipids and complex carbohydrates. To make human milk oligosaccharides (HMOs), the relevant biosynthetic pathway is reconstructed in the host organism: genes encoding specific glycosyltransferases are introduced, and availability of the right nucleotide-sugar donors is tuned, so the microorganism can assemble the target HMO structure starting from lactose and additional simple feedstocks such as glucose or sucrose. The engineered strain is then grown in a controlled bioreactor, where it converts these feedstocks into the target HMO.
coli remains the standard workhorse. Its genetics are familiar, its metabolism is well mapped, and fermentation behaviour is considered predictable.
2’FL synthesis starts with lactose as the acceptor substrate and GDP-fucose as the donor activated sugar. A fucosyltransferase then drives α1→2 fucosylation of lactose.
Yes, a synthetic HMO can match the molecule found in human milk. It has to match on monosaccharide composition, linkage position, anomeric configuration, and full structural assignment. HPLC, LC-MS, and NMR are normally used to prove that identity. For more structural background, see complete HMO guide.
The hard parts are usually enzyme specificity, nucleotide-sugar supply, lactose transport, pathway burden, and side-product formation. At scale, additional challenges emerge around oxygen transfer, feed distribution gradients, CO2 removal, pH control, and the management of process-related impurities in downstream purification.
Theoretically yes, but not all 200-plus structures are within easy reach. The basic neutral, sialylated and fucosylated HMOs are already commercially accessible. Although some very specific structures such as branched (e.g. LNH or LNnH) or multi-decorated (e.g. DSLNT) HMOs remain challenging, many other complex structures (e.g. LNFP, LNDFH, or LST variants) are also already feasible today at small scale, yet remain to be commercialised. The problem is no longer how to make a complex carbohydrate. The challenge is which carbohydrate to make.
Cleaning up the product depends on the host, where the HMO sits, and what impurities come along for the ride. Downstream processing can involve cell removal, clarification, decolourisation, desalting, chromatographic purification, membrane concentration, crystallisation, or spray drying.
HPLC gives the purity picture and related carbohydrate profile. LC-MS helps confirm molecular identity. NMR is the go-to for linkage position and anomeric configuration. Extra release tests may cover residual protein, DNA, endotoxin, moisture, heavy metals, and microbiology.
Costs have come down as strains and processes have improved. Better titre, stronger rate, higher yield, tighter fermentation control, smarter donor-pathway engineering, and more efficient purification all helped. The final economics largely depend on production scale.
Several fermentation-derived HMOs now have regulatory clearance in major food and infant nutrition markets. The status is still molecule-specific and region-specific. Safety, identity, purity, manufacturing, and toxicology data are key topics for each ingredient.
While Inbiose strives to ensure the accuracy and timeliness of the information presented in this document, the company makes no guarantees or assurances regarding the precision, reliability, or completeness of the content. This information is intended solely for scientific and technical purposes in business-to-business communications. Local regulations and requirements should be reviewed when considering product labeling or advertising to end consumers. The information provided does not constitute medical or scientific advice, diagnosis, or treatment, and is offered without any warranties, express or implied. Inbiose shall not be held liable for any damages resulting from the use or interpretation of the material presented herein. The reader assumes full responsibility for their use of the information. No liability is accepted for any infringement of patents resulting from the use of the information provided. Content may be updated without prior notice. For further information, please contact your local Inbiose representative. All trademarks mentioned are owned by or licensed to Inbiose unless otherwise noted.